MHRA-Compliant Preclinical Safety and Toxicology Studies for Advanced Therapy Medicinal Products (ATMPs)

PROVEN INTELLIGENCE ACCELERATING NEXT-GENERATION THERAPIES

MHRA-Compliant Preclinical Safety and Toxicology Studies for Advanced Therapy Medicinal Products (ATMPs)

MHRA-Compliant Preclinical Safety and Toxicology for ATMPs

CELL & GENE | RNA | BIOLOGICS

A robust preclinical safety and toxicology program is the foundation for a successful Clinical Trial Application (CTA) with the UK’s MHRA. For Advanced Therapy Medicinal Products (ATMPs), this requires a scientifically justified, risk-based approach that addresses unique modality-specific challenges such as vector biodistribution, immunogenicity, and long-term expression. Franklin Biolabs executes GxP-compliant studies designed to generate a comprehensive data package that withstands rigorous regulatory scrutiny and supports first-in-human trial designs.

    What are the key components of an MHRA-compliant ATMP toxicology study?

    A: A complete program includes dose-range finding, pivotal GxP toxicology, biodistribution and shedding, immunogenicity (including anti-drug and neutralizing antibodies), and, where relevant, tumorigenicity and vector integration analysis.

    How do you address the unique challenges of AAV vector biodistribution?

    A: We employ validated qPCR and ddPCR methods to quantify vector genome persistence and expression in all relevant target and non-target tissues, providing precise data on vector localization and clearance kinetics.

    What is the typical timeline for a full CTA-enabling toxicology program?

    A: A comprehensive IND/CTA-enabling program, from study design to final report, generally follows an 18-24 month timeline, contingent on the complexity of the ATMP and the specific study designs required.

    How does Franklin Biolabs ensure GxP compliance for UK submissions?

    A: Our programs operate within a GxP framework, managed by a dedicated Quality Assurance unit. All study conduct, data analysis, and reporting strictly adhere to current MHRA guidelines for nonclinical development.

A close-up, detailed shot of a Sartorius Stedim Biotech BIOSTAT STR® single-use bioreactor in a laboratory setting.

Defining the Regulatory Pathway for ATMPs in the UK

The nonclinical development of ATMPs for the UK market demands a deep understanding of MHRA expectations. Unlike conventional biologics, the regulatory guidance for cell and gene therapies necessitates a flexible, science-led strategy. The nonclinical program must be tailored to the specific modality, mechanism of action, and intended clinical application.

The primary objective is to characterize the pharmacologic and toxicologic profile of the investigational product. This involves defining a safe starting dose for clinical trials, identifying potential target organs for toxicity, and evaluating the reversibility of any adverse findings. High-Fidelity In Vivo Intelligence for Complex Modalities.

Core Components of an ATMP Safety Program

A successful CTA submission is built upon a series of well-designed and executed nonclinical studies. Each study addresses a specific question fundamental to understanding the product’s safety profile.

  • Biodistribution and Vector Shedding: These studies determine the distribution, persistence, and clearance of the ATMP from the body. Quantifying vector presence in non-target tissue is a key safety consideration for the MHRA.

  • Immunogenicity Assessment: The potential for an immune response against the vector or transgene product is a significant risk. Validated assays to detect and characterize binding and neutralizing antibodies are a standard program component.

  • Dose-Range Finding and GxP Toxicology: Pilot dose-range finding studies inform the design of the pivotal, GxP-compliant toxicology study. This definitive study establishes the no-observed-adverse-effect-level (NOAEL) required for clinical dose selection.

  • Histology and Bioanalysis: Comprehensive histologic evaluation of a full range of tissues provides insight into microscopic changes. This is correlated with bioanalytical data to build a complete safety picture.

The strategic goal of these integrated studies is to establish a favorable safety profile, a principle demonstrated in AAV gene therapy development programs that have successfully characterized therapeutic benefits alongside a well-understood safety margin (PMID: 35333110).

A scientist in a lab coat and gloves looks through a microscope in a laboratory setting, with a blue color overlay.

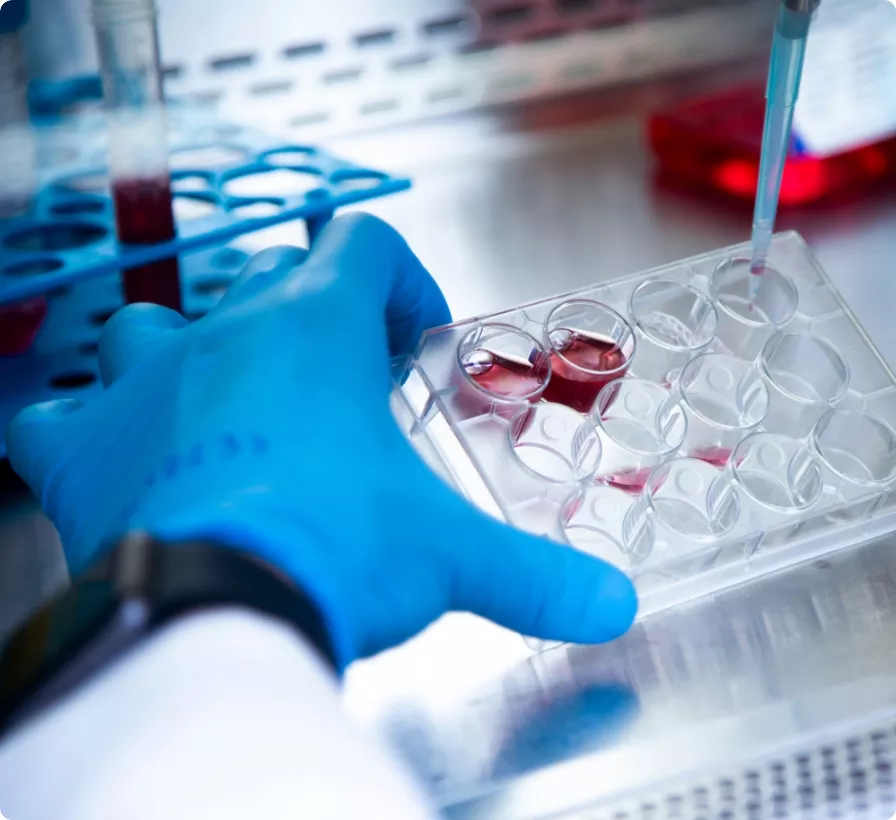
A scientist pipetting a red liquid into a multi-well plate in a laboratory setting.

Animal Welfare and Program Integrity

Scientific validity depends on exemplary animal welfare. Franklin Biolabs is fully committed to the highest ethical standards in all in vivo research, operating in alignment with AAALAC and USDA guidelines. Our animal welfare strategy is centered on the 3Rs:

  • Replacement: Using non-animal methods where scientifically valid.

  • Reduction: Minimizing the number of animals used per study to the amount needed for statistical significance.

  • Refinement: Optimizing all procedures to enhance animal well-being.

From Nonclinical Data to Clinical Trial Application (CTA)

Navigating MHRA requirements with precision toxicology is our focus. The data generated in our >100,000 sq ft GxP-compliant facility is compiled into a submission-ready package designed for regulatory success. Since 2019, programs conducted at our facilities have achieved a 100% IND success rate, a track record of quality we continue under the Franklin Biolabs brand, which launched in 2024. We deliver the robust, defensible data required to confidently advance your ATMP into the clinic.

Scientific Process Diagram

This content is for informational purposes. For guidance specific to your therapeutic program, please contact our team for a consultation.